What does it mean that DEOMED® socks are medical devices? 5 things to know before you buy
Product packaging carries a range of markings, symbols and other information. Some explain what the product is intended for, while others help identify it or provide details about safety and regulatory compliance. DEOMED® socks are marketed as Class I medical devices. What exactly does that mean? Does the CE marking confirm that the product is effective? How do you read the label, and what information can help you make an informed purchase? What should you look for when choosing socks for a specific health need? Here are five things to know before you buy.
1. What does it mean when a product is a medical device?
Some products on the market are described as promoting health, helping to prevent health problems, or meeting the needs of people with specific conditions. These descriptions alone do not give a product medical device status.
Under Regulation (EU) 2017/745 of the European Parliament and of the Council, known as the Medical Device Regulation (MDR), whether a product qualifies as a medical device depends, among other things, on its intended purpose and whether it meets the applicable requirements. In Poland, the Act of 7 April 2022 on Medical Devices applies alongside the MDR.
Many different products intended for medical purposes can qualify as medical devices. Whether a particular product falls within this definition depends on its specific intended purpose and how it qualifies under the applicable rules.
What are the manufacturer’s obligations?
Depending on the type and class of the device, the manufacturer must, among other things:
- define the product’s intended purpose
- assess its safety and performance
- prepare the required technical documentation
- carry out the appropriate conformity assessment procedure
- ensure that the device is correctly labelled and that users receive the necessary information
- monitor the device’s safety after placing it on the market
What does this mean for consumers?
A medical device has a defined intended purpose, and the manufacturer is responsible for ensuring it meets the legal requirements. Users should have access to information about what the product is intended for and how to use it.
2. What does Class I mean for a medical device?
Medical devices are grouped into classes based on their intended purpose and the risks associated with their use. The MDR sets out the classification rules.
Class I covers devices with the lowest level of risk. These devices still require a safety assessment and must meet all applicable MDR requirements.
Who assesses the conformity of a Class I device?
For a standard Class I device, the manufacturer carries out the conformity assessment, prepares the required documentation and draws up the EU declaration of conformity. Certain types of Class I devices are subject to additional procedures involving a notified body.
3. What does the CE marking mean?
The CE marking is one of the most important markings found on medical devices. It indicates that the manufacturer declares the product’s conformity with the requirements of the relevant EU legislation. The manufacturer must carry out a conformity assessment before affixing the CE marking.
The CE marking indicates that the device meets stringent safety standards and legal requirements. It does not serve as a universal certificate of effectiveness, guarantee a cure or indicate that a doctor has individually approved the product.
What else should you check besides the CE marking?
- who manufactures the device
- what the device is intended for
- what the instructions say, including whether there are any contraindications
4. Clinical evaluation and clinical investigation. What’s the difference?
Two similar terms often come up in discussions about medical devices: “clinical evaluation” and “clinical investigation”. They refer to different processes.
- Clinical evaluation is the ongoing analysis of scientific and technical data and information about a device. Its purpose is to determine whether the available evidence supports the claimed performance and confirms the device’s safety.
- Clinical investigation is a planned study involving human participants, such as patients or other study subjects.
A separate clinical investigation involving patients is required only for some medical devices. The clinical evaluation required by the MDR still applies.
5. How do you read medical device packaging?
A medical device label helps you identify the product, find out who is responsible for it and understand how to use it safely. Here are the main things to look for:
- Product name and manufacturer details — identify the entity legally responsible for the device.
- Intended purpose — tells you what the manufacturer designed the product to be used for.
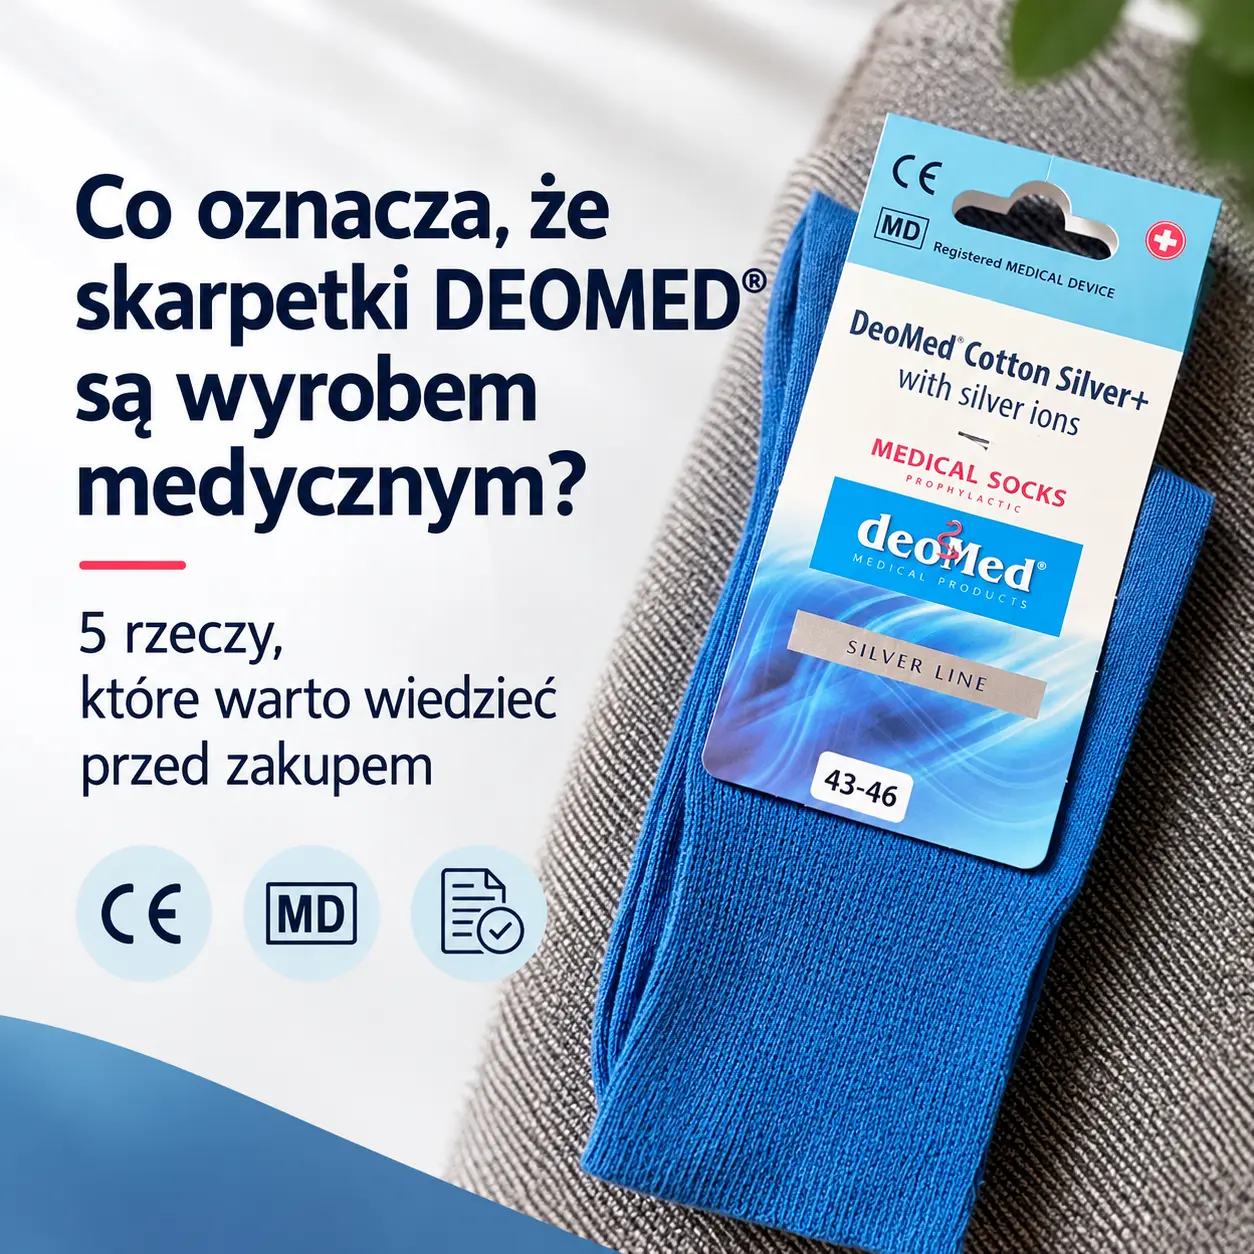
- MD symbol and CE marking — indicate the product’s regulatory status on the EU market.
- Lot number (LOT) or serial number (SN) — helps identify the product when, for example, you make a complaint or report an adverse event.
- Instructions for use and warnings — explain how to use and care for the device and list any contraindications.
What should you keep in mind before buying?
The terms “medical device”, “Class I”, “CE marking” and “clinical evaluation” each refer to a different aspect of the product.
- Medical device — a product with a specific medical purpose that is subject to applicable legal requirements.
- Class I — a classification based on the device’s intended purpose and risk.
- CE marking — a marking that indicates conformity with applicable EU legal requirements.
- Clinical evaluation — an analysis of data on the device’s performance and safety.
When choosing a product for a specific health need, check its intended purpose, labelling and instructions for use alongside its name and marketing description.